Posts

Hidden Markov Models

Hidden Markov Models Hidden Markov Models is one of the popular machines learning approach that can be used to decipher highly complex biological data. One example would be CG-Island. CG is the least frequent nucleotide in many genomes Reason: C within CG is easily methylated Resulting methylation has tendency to mutate into T However methylation is suppressed around genes in areas called CG-Island. Here CG appears relatively frequently. Fair Bet Casino: Through it CG-Islands can be modeled. Dealer flips coin and player bets on outcome: heads or tails Dealer uses either a biased coin (head with probability ¾) or fair coin head or tail equally likely Given a sequence of coin tosses, problem to find out when dealer used a biased coin or a fair coin Obviously, if you observe long line of heads it’s likely that dealer used biased coin and if you observe even distribution of heads and tails he likely used fair one.

P versus NP problem

Image
Millennium Prize Problems are seven problems in mathematics that were stated by Clay Mathematics Institute in 2000 Out of the 7 problems, one (Poincare conjecture)  problem has been solved If some one solves any of the problem, he is awarded with US $1,000,000 prize Poincare conjecture problem was solved by Grigori Perelman on year 2010,  but he declined  to receive the prize. The seven problems are: P versus NP problem Hodge conjecture Poincare conjecture (solved) Riemann hypothesis Yang-Mills existence and mass gap Navier-Stokes existence and smoothness Birch and Swinnertori-Dyer conjecture P versus NP Problem What is P? This class of problem are easy to solve. We can have algorithm to solve this type of problem within a reasonable amount of time. e.g. Check whether a particular number exists in a list. The simplest solution would be "linear search" algorithm, we check each number in turn until we find th...

Synthetic Life: Now possible…..

Now creating an artificial organism that suits our needs comes under realms of possibilities. The well known maverick in the science society named Craig Venter and his colleagues were first to make such an effort and were successful. The entire procedure to reach this pinnacle is explained by Craig Venter in an interview. What I'm going to tell you about in my 18 minutes is how we're about to switch from reading the genetic code to the first stages of beginning to write the code ourselves. It's only 10 years ago this month when we published the first sequence of a free living organism, that of haemophilus influenzae. That took a genome project from 13 years down to four months. We can now do that same genome project in the order of two to eight hours. So in the last decade, a large number of genomes have been added: most human pathogens, a couple of plants, several insects and several mammals, including the human genome. Genomics at this stage of the thinking from a littl...

Questions

What is the program saved in the ROM? Software Hardware Romware Firmware Answer: Read-only memory (usually known by its acronym, ROM) is a class of storage media used in computers and other electronic devices. Because data stored in ROM cannot be modified (at least not very quickly or easily), it is mainly used to distribute firmware (software that is very closely tied to specific hardware, and unlikely to require frequent updates) Ramachandran Plot is symmetric for Glycine Proline Serine Threonine Answer: The Ramachandran plot is the 2d plot of the φ-ψ torsion angles of the protein backbone. It provides a simple view of the conformation of a protein. The φ-ψ angles cluster into distinct regions in the Ramachandran plot where each region corresponds to a particular secondary structure. There are four basic types of Ramachandran plots, depending on the stereo-chemistry of the amino acid: generic (which refers to the 18 non-glycine non-proline amino acids), g...

Ab-Initio Protein Structure Prediction

Image
In this type of protein structure prediction 3-D models are built absolutely from scratch . It makes use of physical principle rather than previously solved structures. Prediction is done with no information about protein except amino acid sequence . The computer simulates physical process and forces that drive proteins into native conformation . About Protein Folding : Each specific sequence of polypeptide yields only single , compact, biologically active fold in native state . This fold generally has many sub-states with minor structural differences between them, but all of these sub-states have the same general fold . In other words, under physiological conditions there appears to be one conformation for a given amino acid sequence that has a significantly lower free energy than any other . So, one can search all possible conformation in random fashion until we get lower energy conformation of native state . But this seems to be impossible : Each peptide group has 3 possible co...

Secondary Structure Prediction

Image
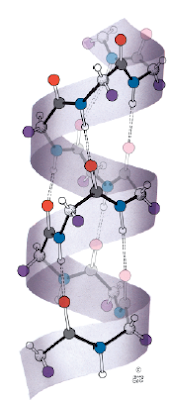
It allows us to find where secondary structural elements ( α helices, β sheets, loops) are located. Secondary Structure Predictors Chou-Fasman COMBINE GORIII PRISM PHD0 PHD3 PSIPred PSIPred uses most recent algorithm that can predict secondary structure of about 80% accuracy. Protein Threading or Protein Fold Recognition Although proteins are of large no, tertiary structural motifs are limited to which most protein belongs. It is speculated that about 1000 distinct protein folding patterns may be present in total. Surprisingly, a few dozen folding patterns account for about half of all known protein structures . This helps to use previously solved structures as starting point. To identify the fold the sequence is compared with all 500+ folds in library of known protein structure . If pair-wise alignment shows less than or nearly 30% identity then it is ideal to be used for protein fold recognitions. When successful, structure from fold recognition may be about 3-6Å RMSD (root mean...

Protein Structure Prediction

Image
Why prediction of protein structure important ? You may ask what is there in protein structure?? That is because structure determines function of the protein. Take the example of enzyme dehydrogenases. It has an NAD-binding site called Rossaman fold ( Dinucleotide binding fold). This fold is made up a pair of βαβαβ subunit. Thus if a protein contain a βαβαβ subunit than it acts as a binding site for a nucleotide . Structure is better conserved than sequences in protein during the course of evolution e.g. take the example of Cytochrome C of eukaryotes and C-cytochromes of prokaryotes in different species(which change with evolution in prokaryotes) where all perform general functions i.e. electron carrier. But different species exibit low degree of similarity of sequence to each other and to that of eukaryotes. They also differ in polypeptide loops on surface . But X-Ray structure are similar particularly chain folds and side chain packing to interior . We require structural knowl...